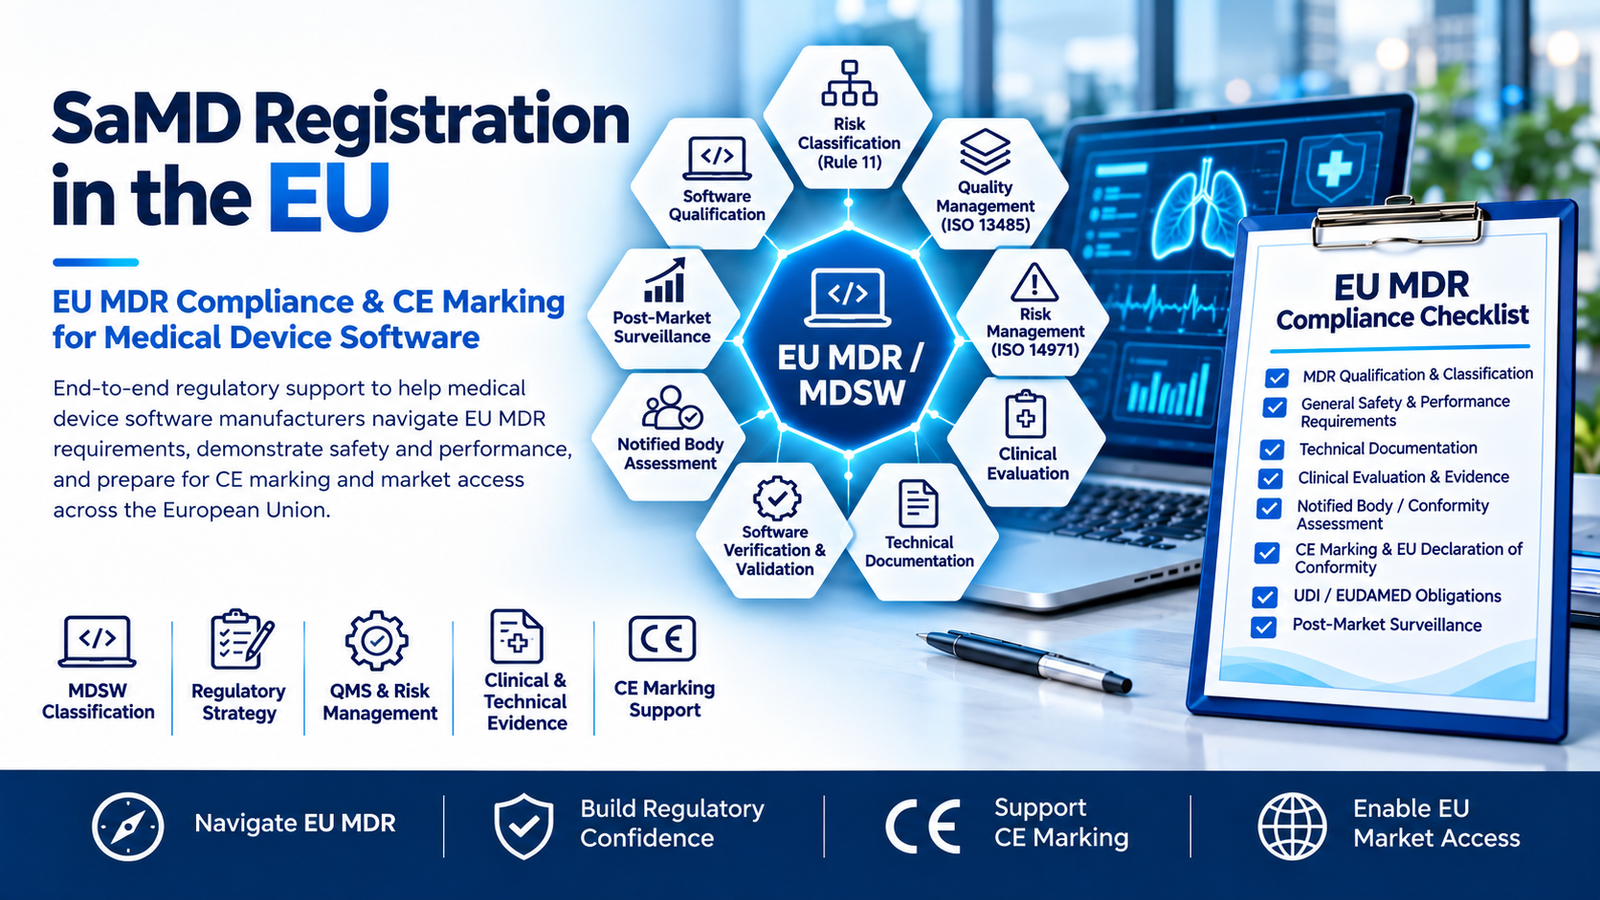
Artificial Intelligence Software as a Medical Device (AI-SaMD) refers to software that utilizes Artificial Intelligence (AI) or Machine Learning (ML) technologies to perform medical functions through software-based processes. These solutions may support diagnosis, disease prediction, clinical decision-making, patient monitoring, treatment planning, medical image analysis, and other healthcare applications.
In the European Union, AI-SaMD products are regulated under the Medical Device Regulation (EU MDR 2017/745). Software that meets the definition of a medical device must comply with applicable regulatory requirements before it can be placed on the EU market and bear the CE marking.
Compliance with EU MDR requirements helps demonstrate that AI-enabled medical software is safe, performs as intended, and maintains an acceptable benefit-risk profile throughout its lifecycle. It also facilitates access to the European market while enhancing confidence among healthcare professionals, patients, and stakeholders.
Support for organizations assessing AI-enabled software under the EU MDR, determining classification, preparing conformity assessment documentation, and managing ongoing regulatory obligations.
Comprehensive support for organizations preparing AI-enabled medical software for EU MDR compliance, CE marking, and ongoing post-market obligations.
Practical, risk-based support for AI-enabled healthcare software seeking EU MDR compliance, CE marking readiness, and sustainable market access.
FAQ
Software is classified according to the requirements of EU MDR, including MDR Rule 11, which specifically addresses software intended to provide information for clinical decision-making.
No. Software intended solely for administrative, lifestyle, wellness, or non-medical purposes may not fall within the scope of EU MDR.
MDR Rule 11 is the classification rule used to determine the risk class of medical software based on its intended purpose and the significance of the information it provides.
Yes. Clinical Evaluation is generally required to demonstrate the safety, performance, and clinical benefit of software regulated under EU MDR.
Yes. Significant modifications to software functionality, intended use, or performance may require additional regulatory assessment and documentation updates.
Yes. Cybersecurity measures should be implemented to protect software integrity, patient information, and overall device performance throughout the lifecycle.
Fill in the form and our team will get back to you shortly.
Reach our regulatory experts directly.